Get in Touch with Guanma

Pharmaceutical label testing assesses the ability of a defined printed label construction to stay attached and legible on its intended container through various forms of handling, exposure and storage. Hence, evidence, apart from an adhesive datasheet or a successful barcode scan, is required for approval. The basis for approval should be traceable to the sample tested, the test conditions, the result measured and acceptance criteria set prior to the test.
Updated October 2026
Quick Specs: define the test before ordering samples
- Test article: facestock, adhesive, print system and protective layer on the intended substrate.
- Method distinction: peel adhesion, curved-container hold, print durability and barcode verification answer different questions.
- Acceptance: use the approved product and process specification; method names supply no universal pass value.
- Evidence: identify the specimen, protocol revision, exposure sequence, result and decision owner.
Pharmaceutical label testing qualifies a printed construction for a defined container and exposure. Evidence for adhesion, legibility and barcode quality needs to be provided separately. Set acceptance criteria before testing and maintain a specimen-bound report.
Key points
- A strong flat-panel peel result leaves curved-vial edge lift untested.
- Cold storage after bonding and application onto a cold, wet surface are different challenges.
- A readable barcode can still have the wrong data or an inadequate quality grade.
- Material qualification supports a defined decision; it doesn’t establish complete drug-label compliance.
What Pharmaceutical Label Testing Should Prove
A pharmaceutical label test assesses a material or printing property under defined conditions. While the scope of the test may include adhesion, retained legibility and barcode quality, approval of the artwork, chemical compatibility and finished-unit labeling remain separate questions when the same pharmaceutical packaging project requires evidence for all of them.
For manufacturing of drugs in the U.S. covered under 21 CFR 211.122(a)-(c), label and packaging materials need to be received under control, representatively sampled, and tested or examined prior to use. Release under such current Good Manufacturing Practice (GMP) controls is dependent on adequate written specifications, and that section doesn’t establish a peel force, wipe count, or grade for barcode for every pharmaceutical label test.
Does pharmaceutical label testing need to be done?
Applicable regulatory requirements and an approved quality system determine which examinations or tests are required. For engineering qualification, a risk-based plan for the intended pack is needed. The acceptable appearance of a sample upon arrival isn’t valid evidence of performance after printing, application and exposure.
Findings in the pharmaceutical industry provide evidence for a defined packaging decision. Patient safety and product integrity also depend on factors outside this material-test matrix; no reductions in labeling errors or product recalls have been measured for this editorial framework. Keep the boundaries visible.
- Material qualification: can the defined construction withstand the intended use?
- Artwork and data approval: does the label contain the approved information?
- Production checks: did the correct label reach the correct pack?
- Compatibility assessment: does the packaging system need separate chemical-suitability evidence?
A 2017 study by Dennis Jenke and coauthors illustrates the last boundary. The abstract made public by the authors describes a model with an LLDPE bottle, a closure, an adhesive label and a foil overpouch, stored for up to 6 months at 40 °C. Label-related substances may migrate into simulated contents without direct label contact. The model migration study abstract doesn’t establish behavior of Guanma stock, every plastic pack or glass vials. Physical durability results alone can’t answer that chemical question.
Approval boundary: Commission the missing assessment when a report answers a different question. Use the pharmaceutical label verification guide, accompanying this document, for approved copy, variable data and release controls.
Define the Label, Container and Packaging Line Before Testing
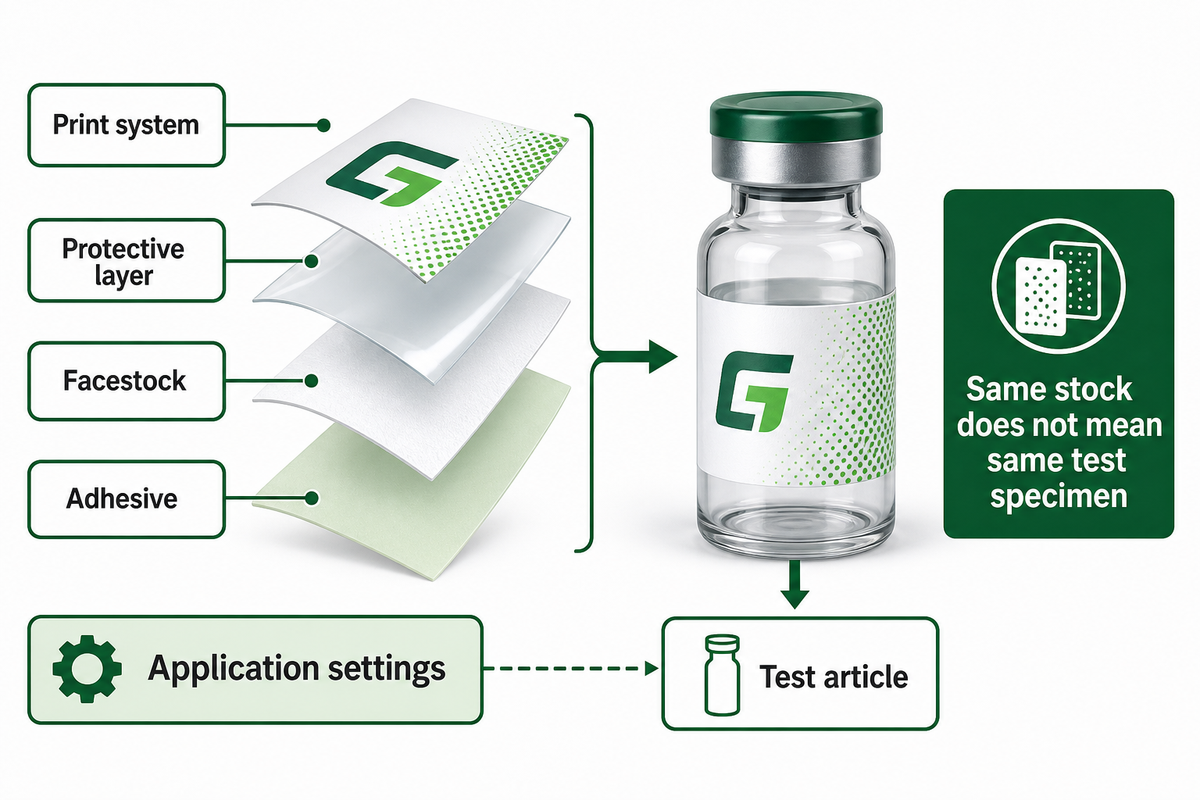
Test results are for a defined label construction and application, as opposed to a facestock name alone. Hold the material grade, print system, container surface and application settings before deciding on samples. Identify intended storage conditions and handling sequence, so that limits of the test may be recognized by others.
Because different facestock stiffness affects how a label follows a curve, and how an adhesive behaves depends on what surface it contacts, the trial description has to include both physical properties and interface conditions, to be able to interpret the results. Ink, ribbon and overlaminate affect how printed information stands up to contact and rubbing. Because of this, a carton panel, coated glass vial and a plastic container require different descriptions, even if their label dimensions are the same. Naming the same stock doesn’t identify the surface or the geometry of the assembly being tested.
“Testing should represent the conditions that the labeling will be exposed to”
3-Stage Label Qualification Plan
The 3-Stage Label Qualification Plan is an editorial sequence to test the construction, challenge the labeled container and check the production line.
- Screen the construction. Select relevant material and print tests using identified stock and controlled preparation.
- Challenge the labeled container. Reproduce surface condition, curvature, exposure and recovery; inspect the complete label.
- Check the pilot line. Assess dispensing, placement and inspection at the intended operating settings before a production decision.
As an example of the challenges of changing existing technology, let’s take the example of changing the paper label on a medicine vial, to a film label. While the peel report for the film is available, the converter wants to use a different ribbon and a different protective coat. The packaging engineer wants to know the proposed printed label construction on the intended vial, including the vial’s surface treatment. Operators then check release from the liner and placement with the chosen applicator. Quality staff inspect the challenged samples against approved criteria and record any deviation. While the peel report for the film is a good screening tool, it doesn’t cover the new print system, container geometry or the dispensing system, and so other tests are needed to validate the changes.
In trials, provide measurements for throughput in containers/min, label dimensions in mm and application temperature in °C. These values are measured inputs to the process. These measured inputs don’t establish that default settings are adequate or constitute recommended settings. A proposal for PET adhesive film still requires this application definition, especially if labeling machines dispense a revised construction using a liner, feed path or application set-up that’s different from the screening test.
Buyer objection: A supplier may provide only raw stock data. Retain the data supplied, but request the converter and packaging-line evidence needed to substantiate the finished construction.
Check Adhesion and Edge Lift on Small Vials
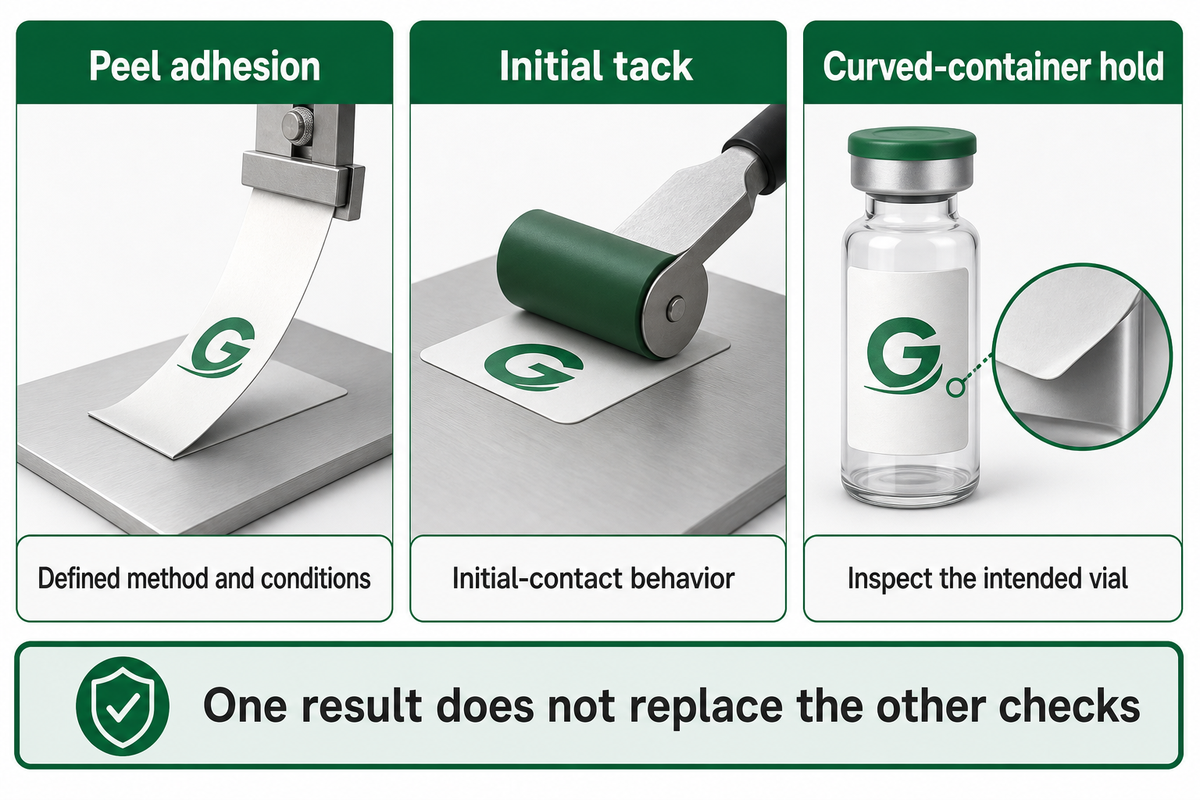
Peel, initial tack and curved-container hold represent various behavioral characteristics. A suitable peel test method should be used to evaluate the behavior of identified samples against one another under comparable conditions, and the labels must be examined on the vials intended for use. Edge lift, tunneling and overlap are behaviors that are dependent upon construction and geometry. A flat-panel number can’t definitely distinguish applicability.
ASTM D6252/D6252M-98(2025) examines pressure-sensitive labelstock peel adhesion at 90°. Scope allowing surfaces of interest and suggestions for the use of representative substrates notwithstanding, stiff or easily stretching backings can complicate judgment of the measured force. A higher measured force does not necessarily mean that the overall label suitability is better. The suitability of the label still requires application-specific evidence beyond this peel method.
| Method | Property or exposure | Limitations / Not suitable for |
|---|---|---|
| FINAT FTM 1 | 180° peel, 300 mm/min | Not interchangeable with a different peel angle or rate. |
| FINAT FTM 2 | 90° peel, 300 mm/min | A flat specimen does not reproduce every vial geometry. |
| ASTM D6252 | 90° labelstock peel | Stiff or stretching backings can distort comparisons. |
| FINAT FTM 8 | Resistance to shear | Not a peel or initial-contact result. |
| FINAT FTM 9 | Loop tack | Not proof of aged container hold. |
| FINAT FTM 13 | Low-temperature adhesion | Define whether the actual application/exposure is represented. |
| FINAT FTM 24 | Mandrel hold | Select applicable geometry; retain a container trial. |
| FINAT FTM 16 / 17 | Spot / immersion chemical resistance | A spot exposure does not reproduce continuous immersion. |
| FINAT FTM 27 | Ink rub for UV-printed labels | Do not silently extend the stated print-system scope. |
Identities of methods come from the FINAT public test-method list and ASTM public scope. They are a selection map, not a complete procedure. FINAT clearly distinguishes its test methods from performance specifications, and the complete procedure should be sought before a test method is performed.
For biological products within its scope, 21 CFR 610.60(e) requires sufficient container area to remain uncovered for its full length or circumference to permit inspection of the contents. Strong adhesion doesn’t address the blocked inspection area, and the clause gives no minimum window width in mm that can be added to a drawing without assessing the container and its applicable conditions.
Comparison rule: review peel angle and rate, substrate, conditioning and dwell before comparing reports. Record edge lift on the intended container, at the agreed location, with the agreed limit. Context for this failure mode is found in the label flagging explanation.
Challenge Cold-Chain Labels With Real Condensation Conditions
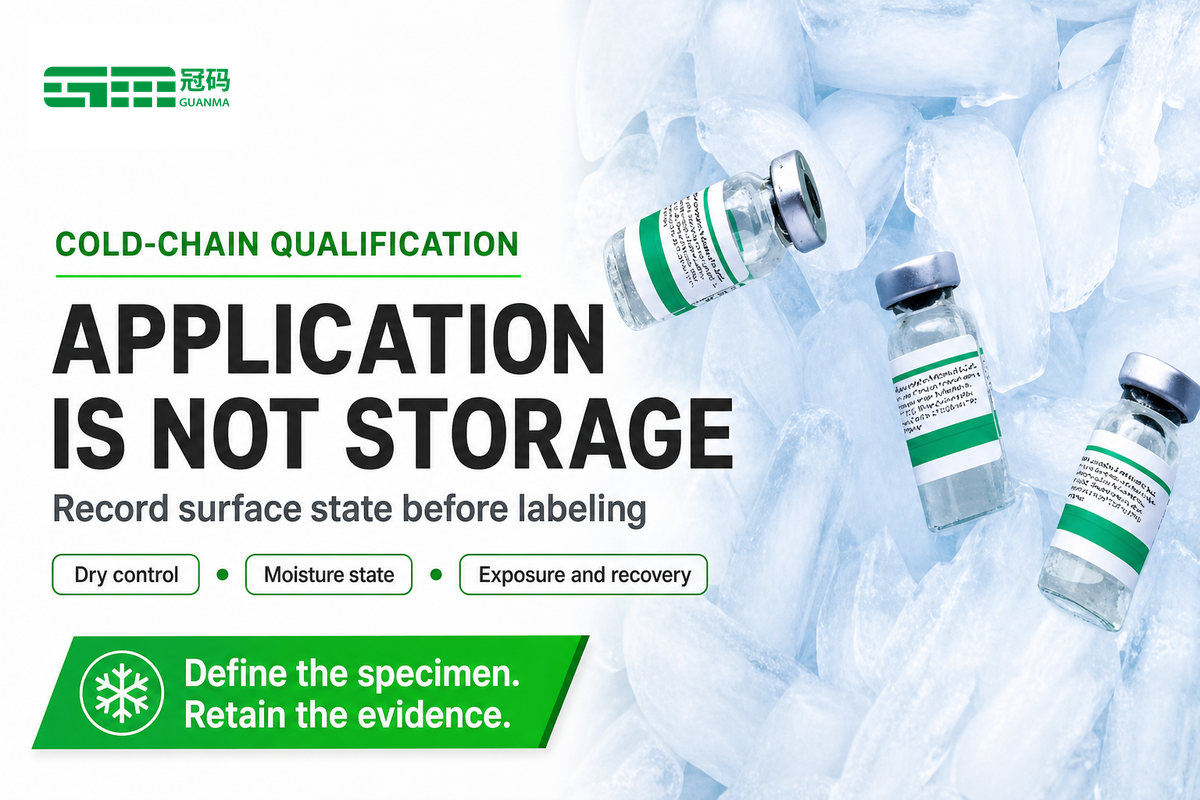
During cold-chain testing of label, the conditions during application must be distinguished from the conditions after the bond has formed. A label applied to a dry container, which has then been placed in a refrigerated condition, hasn’t demonstrated adhesion to a condensation-covered surface. Follow the approved procedures, then inspect the label during exposure and after the prescribed recovery time. The FINAT method list identifies FTM 13 as low temperature adhesion; consult the complete method before using it in a qualification protocol.
Use the product’s approved conditions for the challenge and build the cold-application challenge as part of the study if that’s how the line operates and the study can reproduce the surface state at the point of contact with the label. Include freezing, thawing or sterilizing only if the packaging encounters that. An accelerated challenge should be justified and related to the use; increased challenge severity alone shouldn’t justify the challenge.
- Dry application, later cold exposure: record the initial surface condition and bond-development interval.
- Cold, dry application: verify that preparation and ambient conditions preserve that state.
- Condensation-covered application: document the moisture state and the time between container removal and labeling.
- Recovery inspection: check adhesion, visible defects and legibility at the protocol’s defined inspection points.
Suppose biologic packaging has an approved 2-8 °C storage exposure and its containers are labeled after a controlled transfer into the labeling area. The trial team first documents the container surface temperature and the presence of moisture. If the container is dry, record that condition and compare it with a justified condensation challenge on the material being challenged. Both groups use the same print system with the same application settings. Quality staff document exposure, recovery and observations. These hypothetical trial conditions don’t change the requirements of the approved product. The packaging trial must adhere to those product constraints.
Common mistake: Treating a service temperature claim as proof of cold-surface application is a common mistake. Ask for grade-specific application window and evidence for the tested moisture state. Representative lifecycle exposure, as supported by Susan Ritter, calls for exposure representative of the actual lifecycle; it provides no universal cold dwell or freeze cycle count.
For a 2-8 °C project, the report still needs the measured application temperature, the moisture condition and the exposure duration since recording the storage condition only leaves the bonding step undefined for someone reviewing the evidence later on. Until that sequence is added to the report, the case shouldn’t support this application.
Test Print Against Abrasion and the Actual Wipe Chemical
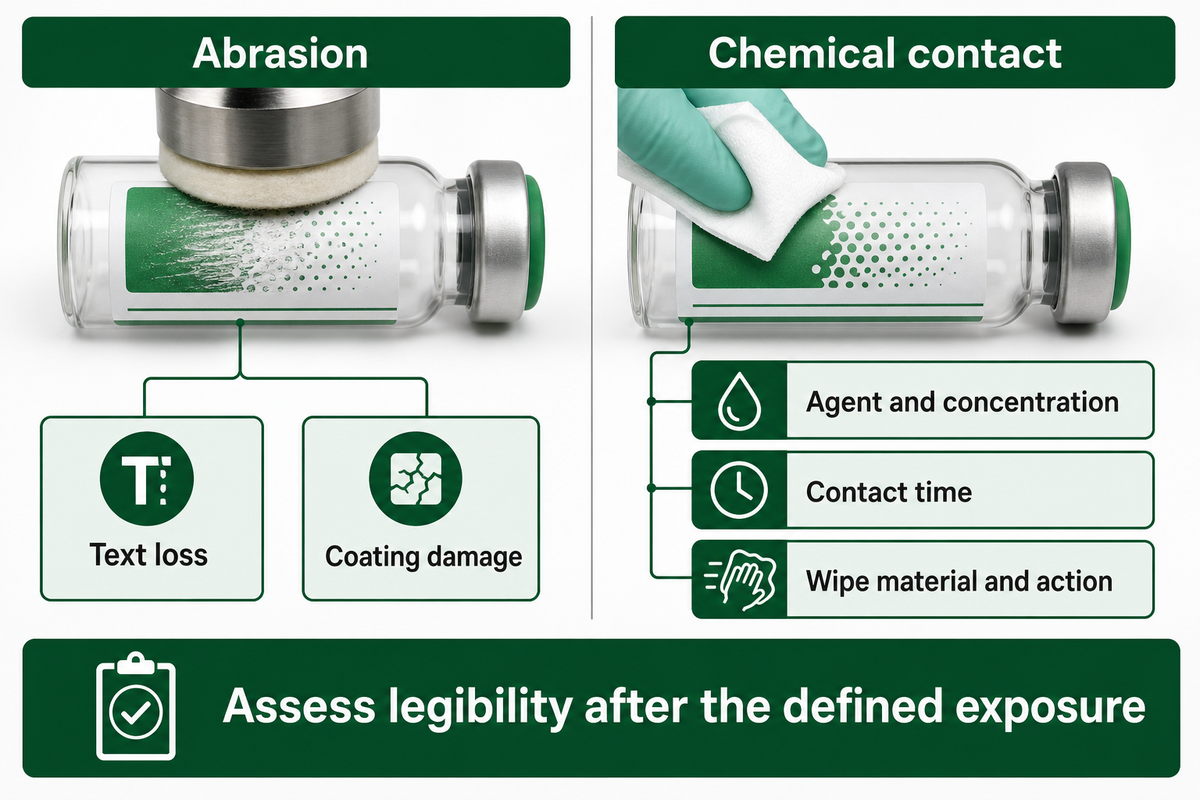
Print durability testing should challenge the complete printed construction, including any contact and/or rubbing that may occur during use. After defining ink, ribbon, coating and facestock, determine the actual reagent, its concentration, contact time, and mechanical action, then evaluate post-exposure quality of the critical information and barcodes, rather than accept an unchanged blank label as evidence.
The public scope of ASTM D5264-98(2019) describes laboratory comparison of printed material abrasion by means of a Sutherland rub tester or similar. Its flat-substrate scope and agreed comparison conditions are the focus of interest. FINAT’s spot and immersion chemical tests address different exposure conditions. Tests for ink adhesion and ink rub are also different test questions.
A wipe trial should define the actual agent by specifying the product or composition, concentration, wetting, contact period, wipe material and the action done to the wipe. The result of this trial should be compared to the approved acceptance criteria for the defined exposure, not to an undefined resistance claim. Choose an endpoint before the test: text loss, smearing, color transfer, protective layer damage, or a reduction in symbol grade. An alcohol-resistant claim doesn’t give any of the above-mentioned conditions by itself. Define the information that a pharmacist and other healthcare professionals need to read after being handled, and then evaluate if it remains legible against the approved reference, not by a general impression that the artwork looks acceptable.
A converter offers a new thermal-transfer ribbon to print variable data on an existing label. The stock and art remain the same. However, the expiry date smears after a representative cleaning exposure. The buyer asks if the stock is defective. The engineer breaks the question into two parts: the ink-adhesion check is with regard to the Print-to-label interface; the wipe test is toward the printed system under that reagent and action; the test report should indicate which question was actually addressed. A controlled comparison would keep the face-stock and printer setting identified, modify the proposed ribbon, and control (save) samples prior and after exposure. The focus is on the system that was exposed to the test condition. Neither a successful dry-rub test nor an untouched drug name would be an acceptable substitute for legible expiry data after contact with the test condition.
A dated example shows why the print system needs its own identity: Steve Katz’s October 2026 DNP ribbon report describes a supplier launch aimed at alcohol-exposed healthcare printing. It reports product claims, not independent proof for another manufacturer’s label construction.
Buyer objection: In the absence of printed samples and the exposure report, request these records to establish whether the supplier tested the printed construction rather than only unprinted stock. If the construction is changed by the use of a chemical-resistant overlaminate, it shouldn’t automatically be considered evidence for every ink and reagent.
Verify Barcode Quality After Exposure
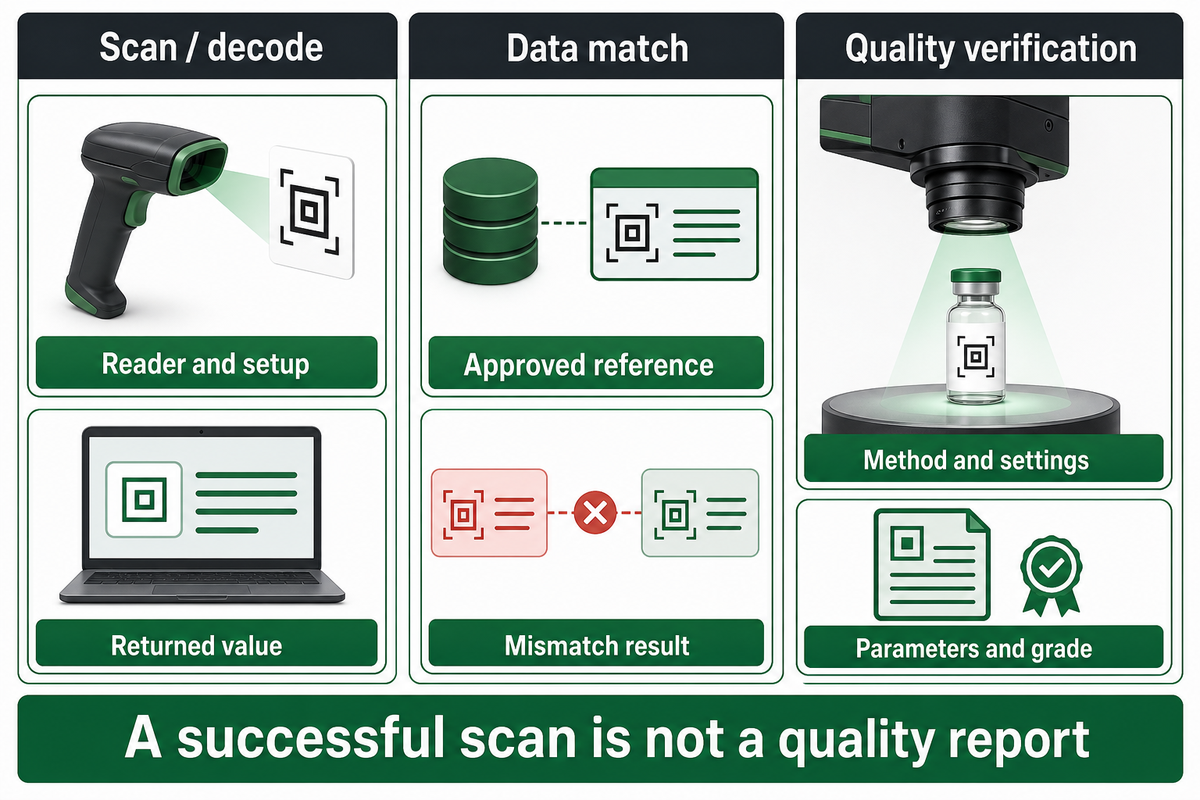
Tests for verifying barcodes require separate records for decoding, approved data and print quality. A test scan shows the decoded information for the barcode in question under the test conditions of the scanner. Because a verifier evaluates a printed symbol against the requirements, some symbols may still be decoded under the test conditions of the scanner, thus repeat the pertinent checks after exposure if the symbol is subject to abrasion, distortion or contamination that may affect the print quality.
Guidance on creating quality-plans for barcode assessment from GS1 identifies ISO/IEC 15416 for linear symbols and ISO/IEC 15415 for two-dimensional symbols. The grade required of symbols depends on the application. While specifying and reporting verifier aperture and wavelength is required, bare letter grades are not sufficient to fully describe the outcome of an assessment.
| Check | Record | Boundary |
|---|---|---|
| Scan / decode | Reader, setup, returned value and specimen | Does not prove every reader can decode. |
| Data match | Approved reference, encoded fields and mismatch result | Correct data do not establish print quality. |
| Quality verification | Applicable method, settings, parameters and grade | A grade does not approve drug content or product identity. |
Can pharmaceutical labels include barcodes or variable data?
Pharmaceutical labels can embed barcodes and variable data as permitted by the approved design and applicable requirements. Other elements such as serial number, lots and expiry date need approved data, content and retained print quality. As such, reference the approved data, record the inspection, and examine human readable information, if any, in the approved design.
For 2D barcodes, like DataMatrix codes, describe the use case and scanning environment of the symbol. GS1’s retail 2D verification explanation clarifies the distinction between scanner and verifier. Their Retail Acceptance Tables aren’t universal pharmaceutical limits; unique identifiers and anti-counterfeiting features need their own approved data rules.
Report decision: Measurement settings are requested when two different grades are given in separate reports. Keep pre-exposure and post-exposure results for same specimen identities. Software inspection records should indicate what has been checked. A beep can’t demonstrate the entire evidence chain; therefore, automation and a beep can’t be considered exhaustive.
Use a 7-Check Label Test Matrix and Record the Decision
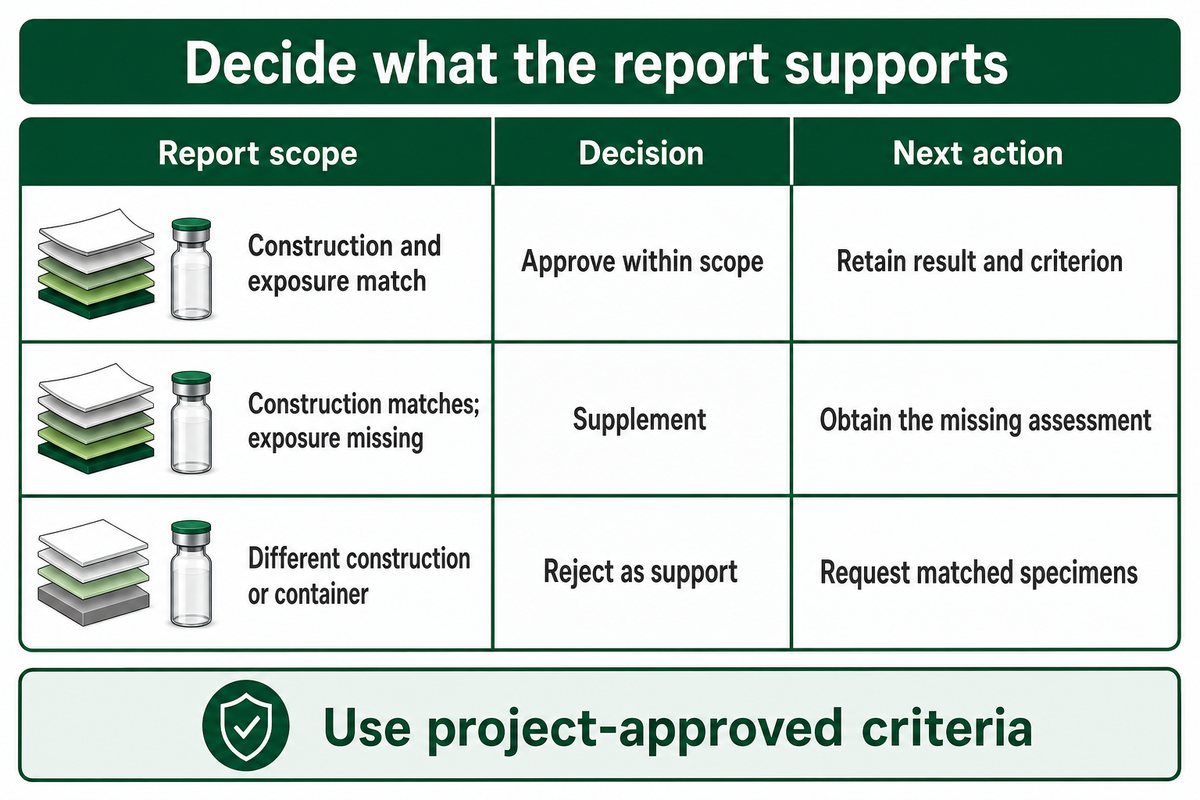
An effective qualification matrix associates each failure mode with a test method, controlled variables, an observed result and approved acceptance criteria. Decide responsibility before testing. Classify each report as applicable, needing supplementation or out-of-scope. Don’t treat every marked pass as interchangeable evidence for the proposed package or a variant that’s being qualified.
7-Check Label Test Matrix
The 7-check Label Test Matrix is an editorial planning tool to link fail modes of printed and processed material to the evidence required for making a defined qualification decision.
| Check | Define before testing | Measure / retain | Limitations / Not suitable for |
|---|---|---|---|
| Construction and specimen identity | Stock, adhesive, print, coating, revision and container | Sample code, batch reference and photographs | Unidentified samples cannot support the proposed construction. |
| Adhesion and initial contact | Applicable peel/tack method, substrate, preparation and dwell | Force with unit, method conditions and failure mode | Do not compare 90° and 180° values as equivalent results. |
| Curved-container hold | Diameter, label dimensions, surface and overlap geometry | Edge-lift measurement, tunneling and inspection images | Flat-panel data leave actual-vial geometry untested. |
| Environmental exposure | Application state, temperature, humidity and lifecycle sequence | Exposure log and defined recovery observations | Cold service alone does not establish wet-surface application. |
| Abrasion and handling | Printed construction, rubbing setup and agreed severity | Critical-text condition, coating damage and symbol result | A dry-rub result does not establish chemical resistance. |
| Chemical contact | Reagent, concentration, contact and wiping action | Smearing, transfer, detachment and retained legibility | One tested reagent does not establish universal resistance. |
| Barcode and data retention | Symbol, approved data, applicable grade and verifier settings | Decode, data match and pre/post verification report | Material tests do not replace drug-label approval. |
Framework basis: The basis for the framework is public FINAT and ASTM methodology scopes, GS1 verification guidance and Ritter’s lifecycle qualification approach. The seven check count is editorial organization. Add other product-related chemical compatibility, tamper evidence or inspection window assessments, outside the scope of this material test matrix.
21 CFR 211.160 requires that appropriate specifications, sampling plans and procedures, along with changes to these, be reviewed and approved by the quality control unit, within the scope of that regulation. Because the samples must be representative and adequately identified, sample numbers and acceptance criteria should be determined by that controlled process; a sample count isn’t assigned by the matrix.
Specimen-to-Report Evidence Sheet
The Specimen-to-Report Evidence Sheet is a handoff record that enables a reviewer to correlate a test result to the actual construction, container and decision it represents.
| Field | Entry to request | Decision supported | Limitations / Not suitable for |
|---|---|---|---|
| Sample identity | Unique sample code and receipt reference | Connect report to physical specimens | A stock-family name is insufficient. |
| Material construction | Facestock, adhesive, coating and grade revision | Check construction match | A changed layer can invalidate the comparison. |
| Print system | Ink/ribbon, printer and controlled settings | Assess printed-system relevance | Blank-stock evidence does not cover printed data. |
| Container | Material, surface treatment, diameter in mm | Check substrate and geometry | Steel-panel evidence may need a vial trial. |
| Application | Surface temperature in °C, moisture and applicator settings | Check bond-formation conditions | Later storage does not define application state. |
| Protocol | Method/version, preparation, dwell and approved deviations | Compare test execution | Different angles/rates need interpretation. |
| Exposure | Actual concentration, duration and sequence | Assess lifecycle coverage | Unspecified severity is not reproducible. |
| Results | Individual observations, units, images and failure modes | Review measured endpoint | A pass stamp hides the sampling basis. |
| Barcode record | Symbol, verifier, aperture, wavelength and grade | Interpret quality measurement | Decode alone supplies no quality grade. |
| Approval | Criterion revision, reviewer, date and limitation | Authorize only the documented scope | Sample approval does not establish every later lot. |
When not to approve the proposed construction
While a report describes the behavior of a system, a material decision should be held when it covers a different adhesive, print system or container. A matched specimen set and a controlled comparison should be requested. A certificate of analysis may define an agreed lot check, but it can’t prove a lifecycle property that was never tested. Separate material certifications from evidence of the performance of the printed label. For an audit, retain the electronic records and associated specimen images in a controlled repository with the protocol revision and decision so a reviewer can trace the reported result without treating the storage location as proof of validity.
Supplement when construction matches but critical exposure is missing. A cold-storage result leaves condensation-covered application unresolved. A barcode decode leaves formal quality verification unresolved. A report from an accredited lab still needs the correct method and scope; Greg Hrinya’s 2024 AD Procerta report is about Avery Dennison’s service and supplies no accreditation claim for Guanma.
Finished unit controls should be active after qualification. Correct-label checks during finishing, and representative completed-unit visual examination and batch production or control records are separately addressed in 21 CFR 211.134. A material report can’t attest to those checks. The QC staff should record unresolved items, the responsible individual, and what evidence is needed to make a release decision. Additional evidence should be provided especially if sample approval is offered as evidence for a finished products release with different construction or exposure history.
Key takeaway
Approve evidence for the construction and conditions it covers. Supplement a missing exposure or measurement. Reject a report as support for a different specimen; a test-method name cannot repair that mismatch.
Send Pharmaceutical Companies a Traceable Sample and Change-Control Brief
Suppliers should define the specimen for pharmaceutical companies, stock manufacturers and converters the same way. Identify who provides the material, who prints it, and who approves it. Agree how construction or process changes will be communicated and how and when evidence should be provided to support those changes, and identify the specific requirements that a matched supplier report must address before it supports the decision.
21 CFR 211.184 requires shipment identity and quantity, supplier name, supplier lot if known, receipt, test results and conclusions, in its records. A good procurement handoff captures these identities for traceability, while keeping the records mandated by the regulation distinct from the editorial sheet this blog is focused on. Use sample to report fields to validate the evidence match, and compare candidate materials, while retaining regulated approval decisions with the authorized quality function.
Guanma declares itself as a manufacturer of self-adhesive paper and film label materials. Its pharmaceutical labelstock offering includes PET and PP synthetic paper and coated paper. Send the container substrate, dimensions, printing process, line speed, and exposure, when requesting candidate samples. These statements don’t create an accredited pharmaceutical laboratory or approval of a specific drug pack.
Send
- Identify stock and print revisions.
- Provide container and exposure details.
- Request sample-bound results and change notification.
Resolve before approval
- Separate supplier and converter responsibilities.
- Confirm who owns each test report.
- Assign quality approval to the authorized role.
Handoff decision: Evidence is compared; engineering defines test article, and quality approves scope. A request to produce compliant labels requires the defined requirements and those functions and roles, not an assurance by the supplier.
Keep Barcode Change Dates Separate From Test Approval
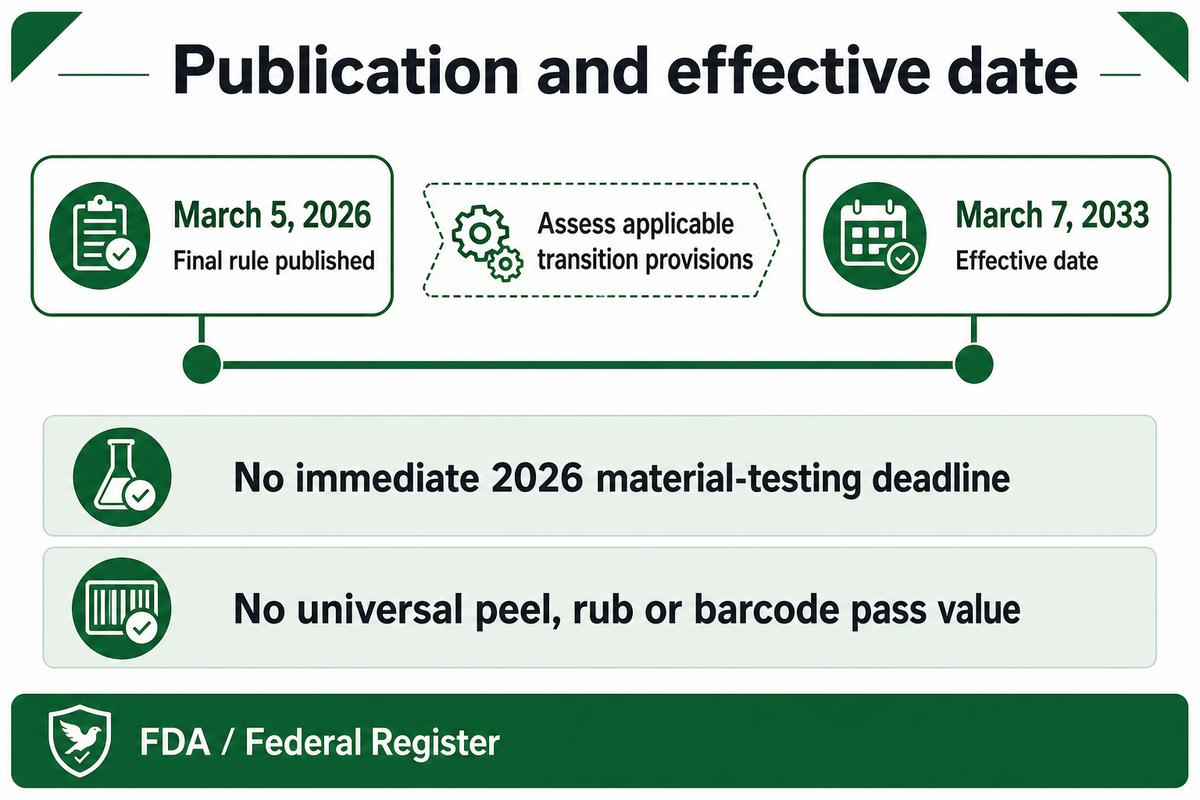
A future identifier change should enter packaging change control with its own dates and impact assessment. FDA published an NDC and drug-label barcode final rule in March 2026, but its effective date is March 7, 2033. The change does not create an immediate 2026 material-testing deadline or a universal barcode grade, so buyers should separate FDA’s implementation milestones from the test conditions used to evaluate the current label construction.
The March 5, 2026 final rule implements a uniform 12-digit format for the FDA assigned national drug code. FDA’s NDC format summary indicates that the effective date is March 7, 2033, and clarifies FDA assigned codes from formats used for other purposes. Take relevant transition provisions for the actual product and labeling program; publication and implementation are different milestones.
For future changes involving the layout or data content of artwork, assess the impact of changes to the placement, size, or print settings of encoded data. More content may not always fit in the same space with the same measured quality. Keep the approved reference and identify the print file and qualify the symbol with respect to its intended packaging. Don’t take for granted an unaltered adhesive means the entire label change is irrelevant.
- Record the applicable regulatory milestone and responsible regulatory reviewer.
- Identify the affected artwork, encoded fields and electronic system interfaces.
- Assess whether printing or application changes require additional qualification.
- Retain the approved revision and the evidence used for the change decision.
Planned boundary: The rule covers qualifying barcode formats and product identifiers. It doesn’t cover a new peel or abrasion pass value. DSCSA product identifier requirements, and other labeling duties, warrant a separate applicability review. A regulatory submission or change record shouldn’t rely on this material-test matrix to show satisfaction of those requirements.
Pharma buyers can prepare a controlled change list without discarding usable evidence because a rule was published. Re-assess the affected construction, printing and data elements when the proposed change is defined. Keep unrelated material results and their original scope and version.
Frequently Asked Questions
How does pharmaceutical label testing work?
Pharmaceutical label testing compares identified specimens with approved criteria under defined application and exposure conditions.
Start by documenting the stock, adhesive, print system and container. Choose methods that address the expected failure modes, then record preparation, exposure and inspection conditions. Compare measured results with the approved specification and retain the supporting images and report. A single test does not cover every property, so missing evidence should be identified before the qualification decision is made.
Can you provide batch records and material certifications?
Ask Guanma which lot documents and material declarations are available for the exact grade being sampled.
State the required document, material code, revision and supplier batch reference in the request. A material declaration, certificate of analysis and drug manufacturer’s batch record serve different purposes. Confirm the issuer and scope of each document rather than assuming the stock supplier owns all production records. Printed-label trial results also need the converter’s print identity and the tested container; a generic certificate cannot supply those missing details.
What materials do you offer for cold-chain pharmaceutical products?
Guanma lists paper and film labelstock options; cold-chain suitability must be established for the selected grade and application.
Send the container material, geometry, application temperature, moisture state and approved exposure sequence when requesting samples. The published pharmaceutical portfolio includes PET, PP synthetic paper and coated paper, but a facestock family alone does not establish cold performance. Adhesive selection and the printing system matter. Require evidence for the proposed construction, and distinguish cold storage after bonding from application onto a cold or wet surface.
Can you produce labels that meet FDA 21 CFR pharmaceutical labeling requirements?
A labelstock sample or material test report cannot establish complete FDA compliance for a finished pharmaceutical label.
Guanma supplies label materials, while the converter and drug manufacturer have separate responsibilities for printing, approved content, controls and release. The applicable product type and market determine the labeling requirements. U.S. rules in 21 CFR Part 211 address manufacturing controls; 21 CFR Part 201 concerns drug labeling and is not an adhesive-material approval scheme. Other product-specific provisions may also apply.
Give the supplier the approved material and performance requirements, then evaluate matched samples and supporting documents. The authorized quality and regulatory functions must determine whether the evidence meets the actual compliance requirements under the approved product program, including the information labels must contain and the evidence needed to assess that information after the intended exposure. No blanket FDA-approved-labelstock claim follows from passing peel, wipe or barcode testing. A prescription drug, OTC product, clinical trial pack or nutraceutical should not inherit another product category’s requirements simply because similar labelstock is used. The same caution applies when a supplier markets pharmaceutical and nutraceutical applications together.
What is your minimum order quantity for pharmaceutical labels?
Request a grade-specific quotation from Guanma; this testing guide establishes no universal minimum order quantity.
Specify material construction, roll width, liner, quantity and sample needs. Separate the raw-stock quotation from the converter’s printed-label order. Confirm current availability and terms directly before committing to production.
What are common printing techniques for pharmaceutical labels?
Pharmaceutical label printing can combine fixed artwork with a separate variable-data printing process.
Flexographic printing, digital printing technology and thermal-transfer variable printing are examples. Qualify the proposed ink or ribbon with its facestock and protective layer. A change in print method can change durability and barcode quality even when the label dimensions and approved artwork remain the same.
Do you offer tamper-evident labels for pharmaceutical products?
Ask Guanma about the exact tamper-evident construction and obtain evidence for its intended pack.
Tamper-evident labels and tamper-evident seals need a defined opening indicator and application trial. A holographic feature alone does not demonstrate tamper evidence. Neither this guide nor a general material listing establishes a product-specific offer, security result or applicable regulatory mandate.
Request samples against your test brief
Send Guanma your container, label dimensions, print system, application conditions and intended exposure. Request candidate labelstock and the grade-specific evidence needed for your qualification plan.
Technical scope: the matrices are editorial planning aids based on public regulations, summaries of methods, and attributed engineering and scientific guidance. They aren’t test results, complete standard procedures, or product approvals. Guanma hasn’t provided proprietary pharmaceutical trial data for this article. The drug manufacturer’s authorized functions are responsible for ensuring the approved requirements and supporting evidence for the product.
Related Articles
- Label durability testing standards and method scope
- Sample qualification and batch acceptance
- PET versus BOPP film label selection
- Guanma label-material manufacturing profile
References & Sources
- 21 CFR 211.122: labeling and packaging materials, eCFR.
- Jenke et al., simulated leaching study, 2017, PDA Journal of Pharmaceutical Science and Technology; public abstract indexed by PubMed.
- Susan Ritter, Planning and Executing Label Qualification, 2020, Packaging Strategies.
- ASTM D6252/D6252M-98(2025), ASTM International; public scope.
- FINAT Test Methods, FINAT; public method list.
- 21 CFR 610.60: biological-product container labels, eCFR.
- ASTM D5264-98(2019), ASTM International; public scope.
- DNP thermal-transfer ribbon launch, October 2026, Steve Katz, Label & Narrow Web; supplier announcement.
- Ten steps to barcode your product, GS1; quality-plan section.
- 2D Barcodes at Retail Point-of-Sale, GS1; general verification distinction only.
- 21 CFR 211.160: laboratory controls, eCFR.
- AD Procerta accreditation report, May 2024, Greg Hrinya, Label & Narrow Web; Avery Dennison service announcement.
- 21 CFR 211.134: drug product inspection, eCFR.
- 21 CFR 211.184: labeling records, eCFR.
- Revising the National Drug Code Format and Drug Label Barcode Requirements, March 5 2026, U.S. Food and Drug Administration, Federal Register.
- National Drug Code Format, U.S. Food and Drug Administration.